Trimetazidine dihydrochloride FAO inhibitor reduces non-cardiovascular mortality in ischemic heart disease breast cancer patients
Copy code to add this image to your page
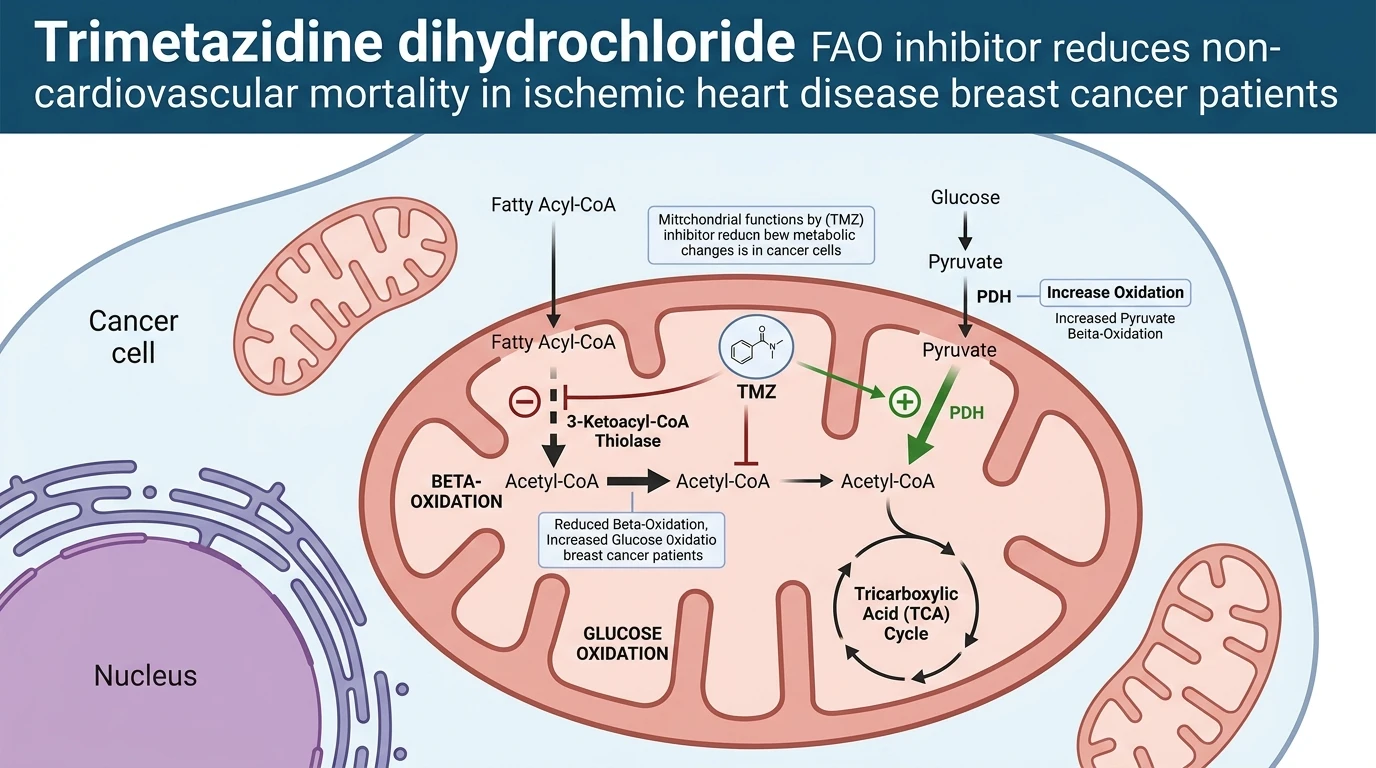
Mitochondrial energy metabolism dictates cellular survival under hypoxic and nutrient-restricted microenvironments. Malignant breast tumors frequently exhibit metabolic plasticity, upregulating fatty acid beta-oxidation to fuel proliferation, membrane synthesis, and bioenergetic adaptation. Trimetazidine dihydrochloride operates as a selective inhibitor of mitochondrial 3-ketoacyl-CoA thiolase, the terminal enzyme in the fatty acid oxidation cascade. By impeding mitochondrial beta-oxidation, the Trimetazidine dihydrochloride FAO inhibitor stimulates pyruvate dehydrogenase complex activity. This enzymatic shift reroutes acetyl-CoA production toward pyruvate-derived glucose oxidation, optimizing cellular oxygen efficiency and altering intracellular metabolite pools.
In a retrospective clinical cohort comprising 2,526 individuals diagnosed with concurrent ischemic heart disease and breast cancer, investigators systematically examined the long-term survival impact of metabolic modulation. Patient records were analyzed to evaluate all-cause and cause-specific mortality differences between individuals receiving trimetazidine dihydrochloride and those prescribed long-acting nitrates as active controls. Within the subcohort of individuals with pre-existing breast cancer, trimetazidine administration demonstrated an independent association with reduced all-cause mortality (adjusted hazard ratio 0.48, 95% confidence interval 0.48–0.98, P = 0.04).
This observed survival advantage was driven principally by a marked reduction in non-cardiovascular mortality rather than ischemic event mitigation alone. Extended sensitivity analyses incorporating both pre-existing and de novo breast cancer cases confirmed a persistent survival benefit (adjusted hazard ratio 0.75, 95% confidence interval 0.60–0.94, P = 0.01). These epidemiological findings suggest that anti-ischemic metabolic agents can exert off-target antitumor activity in human patient populations.
At the biochemical level, cancer cells utilize high rates of fatty acid oxidation to generate NADPH and ATP, mitigating oxidative stress and sustaining biomass synthesis. Inhibition of 3-ketoacyl-CoA thiolase disrupts this lipid reliance. By forcing a substrate transition toward glucose oxidation, pyruvate dehydrogenase flux increases. Pyruvate dehydrogenase converts pyruvate into acetyl-CoA with higher stoichiometry of ATP produced per mole of consumed oxygen. In neoplastic tissues, this metabolic shift compromises the bioenergetic flexibility required for tumor survival under oxidative stress.
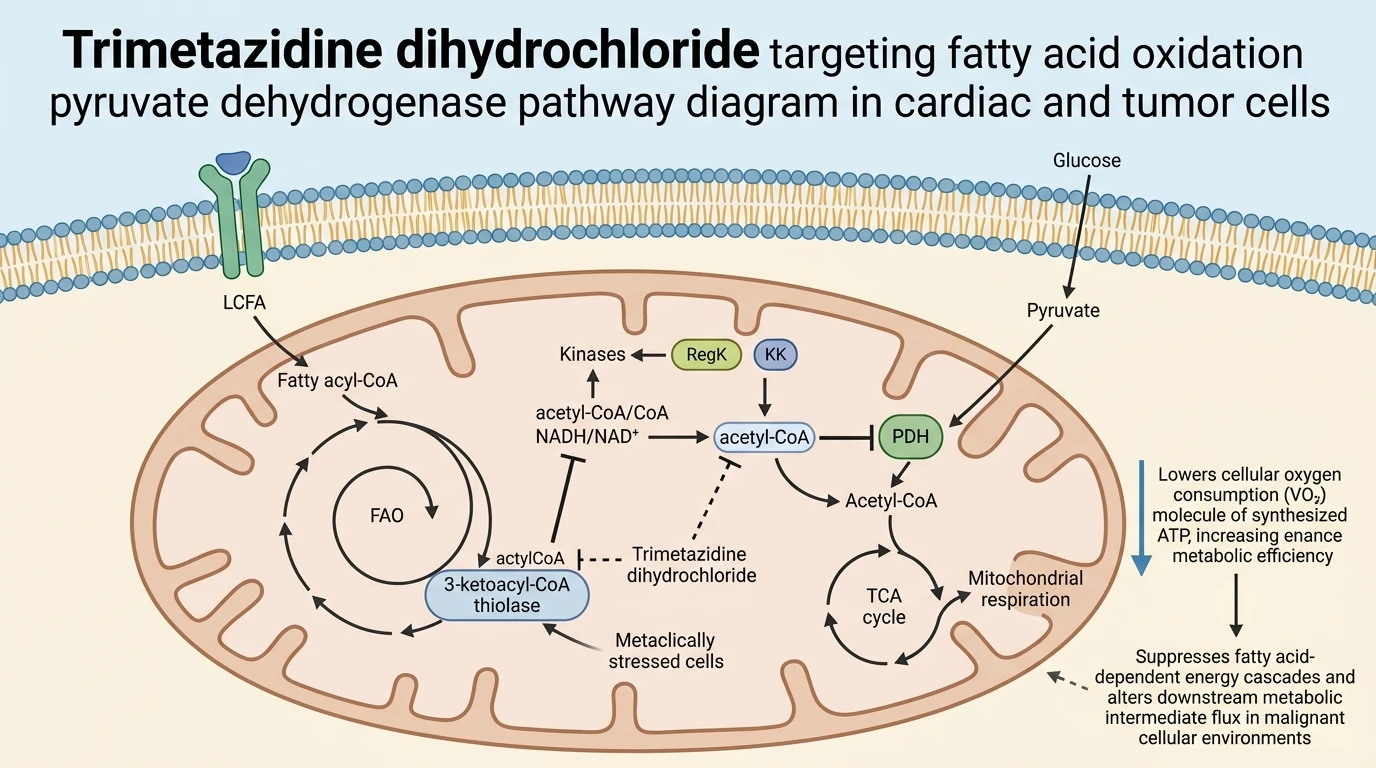
The biochemical interplay between lipid and carbohydrate oxidation is historically defined by the Randle cycle. In this cycle, elevated fatty acid oxidation generates an accumulation of mitochondrial acetyl-CoA and NADH, which allosterically activates pyruvate dehydrogenase kinase. Activated kinase phosphorylates and inactivates the pyruvate dehydrogenase E1 alpha subunit, suppressing glucose oxidation. By inhibiting mitochondrial 3-ketoacyl-CoA thiolase, trimetazidine reverses this cascade. The resulting drop in the acetyl-CoA/CoA ratio restores unphosphorylated, functional pyruvate dehydrogenase. This enzymatic restoration reactivates mitochondrial pyruvate flux, driving efficient glucose-derived oxidative phosphorylation while depriving tumor cells of fatty acid-derived intermediates required for membrane lipid biosynthesis.
The distinction between active control cohorts remains critical when interpreting observational pharmacoepidemiology. Long-acting nitrates alter vascular hemodynamics without directly shifting intracellular substrate utilization. Trimetazidine, conversely, reconfigures mitochondrial bioenergetics at therapeutic plasma concentrations. The observed reduction in non-cardiovascular deaths among breast cancer patients suggests that metabolic reprogramming via pyruvate dehydrogenase activation may limit oncogenic expansion or treatment resistance.
Systemic administration of trimetazidine alters intracellular enzymatic kinetics without producing overt systemic toxicity. Mitochondrial fatty acid oxidation suppression reduces cellular dependence on exogenous lipid uptake. Consequently, tumor microenvironments characterized by limited glucose availability become metabolically hostile to cancer cells forced into pyruvate oxidation. This phenomenon highlights how targeted metabolic modulation alters tumor cell viability independently of conventional cytotoxic pathways.
Although retrospective observational data cannot definitively establish direct causality, the statistical independence of the hazard ratios highlights a meaningful pharmacodynamic response. These clinical observations align with pre-clinical cancer models where metabolic substrate switching induces energy crisis in lipid-dependent malignant cells. The combination of clinical survival metrics and defined mitochondrial mechanisms position trimetazidine dihydrochloride as a candidate for targeted oncology repurposing strategies. Further prospective clinical trials remain necessary to establish optimal dosing, duration, and patient stratification biomarkers.
Epidemiological Outcomes and Metabolic Reprogramming Mechanisms
Copy code to add this image to your page
Trimetazidine dihydrochloride selectively inhibits mitochondrial 3-ketoacyl-CoA thiolase, suppressing fatty acid oxidation and activating pyruvate dehydrogenase. This enzymatic shift redirects metabolic flux from beta-oxidation toward glucose oxidation, increasing ATP production efficiency per oxygen molecule consumed. In breast cancer cells, forcing this substrate transition disrupts metabolic flexibility, lowers NADPH generation, and increases sensitivity to metabolic stress. Retrospective clinical data demonstrate that this pharmacological metabolic reprogramming correlates with significantly reduced non-cardiovascular mortality in ischemic heart disease patients with co-occurring breast cancer.
Reduction of Non-Cardiovascular Mortality in Clinical Cohorts
Analysis of the 2,526-patient cohort revealed a distinct survival advantage among individuals receiving trimetazidine compared to active nitrate controls. The adjusted hazard ratio of 0.48 in pre-existing breast cancer cases indicates a notable reduction in overall mortality risk. Cause-specific mortality decomposition revealed that this survival benefit stemmed primarily from decreased non-cardiovascular deaths. Statistical adjustments accounted for age, cardiovascular comorbidities, baseline oncological staging, and concurrent systemic cancer therapies. Sensitivity analyses encompassing de novo breast cancer cases yielded an adjusted hazard ratio of 0.75 (95% CI 0.60–0.94, P = 0.01). The consistency across subgroups strengthens the association between metabolic intervention and patient survival. These epidemiological patterns suggest a systemic antitumor effect in patients receiving the Trimetazidine dihydrochloride FAO inhibitor.
Mitochondrial Substrate Switching Compromises Tumor Energy Advantage
Mitochondrial 3-ketoacyl-CoA thiolase inhibition suppresses long-chain fatty acid beta-oxidation within cardiac and neoplastic tissues. Reduced fatty acid oxidation relieves acetyl-CoA-mediated feedback inhibition on the pyruvate dehydrogenase complex. Enhanced pyruvate dehydrogenase activity accelerates pyruvate entry into the tricarboxylic acid cycle. Glucose oxidation requires significantly less oxygen per ATP molecule generated than beta-oxidation. In lipid-reliant breast cancer subtypes, this substrate shift disrupts redox homeostasis and depletes cytosolic NADPH pools. Cellular ATP production efficiency drops relative to basal metabolic demands under hypoxic stress. The resulting bioenergetic disruption restricts tumor adaptation without compromising non-malignant tissue survival. Trimetazidine dihydrochloride thus modulates cellular energy dynamics through targeted enzymatic inhibition.
Translational Implications for Metabolic Repurposing in Oncology
Repurposing established cardiovascular therapeutics offers an accelerated pathway for clinical oncological interventions. Trimetazidine possesses a well-characterized safety profile across decades of clinical cardiology administration. Its capacity to alter pyruvate dehydrogenase flux without inducing systemic hypoglycemia provides a favorable pharmacodynamic window. Current observational metrics indicate reduced non-cardiovascular death rates, yet prospective randomized controlled trials are required to validate efficacy in non-cardiac cancer populations. Molecular biomarker profiling, including baseline lipid dependence and pyruvate dehydrogenase phosphorylation status, will be critical for patient selection. Trimetazidine dihydrochloride FAO inhibitor therapy represents a pragmatically viable metabolic adjuvant strategy in breast cancer management.